Key Points
- QURE shares retreat 2.98% to $24.09 despite receiving FDA Priority Review status for ifezuntirgene inilparvovec.
- Regulatory agency establishes May 1, 2027, as target action date under Accelerated Approval pathway.
- Company intends to initiate screening for confirmatory study enrolling 200 Huntington’s patients before 2026 ends.
- Investigational therapy AMT-130 leverages proprietary miQURE gene-silencing technology platform.
- Stock breaks through $24.83 technical support level despite achieving significant regulatory progress.
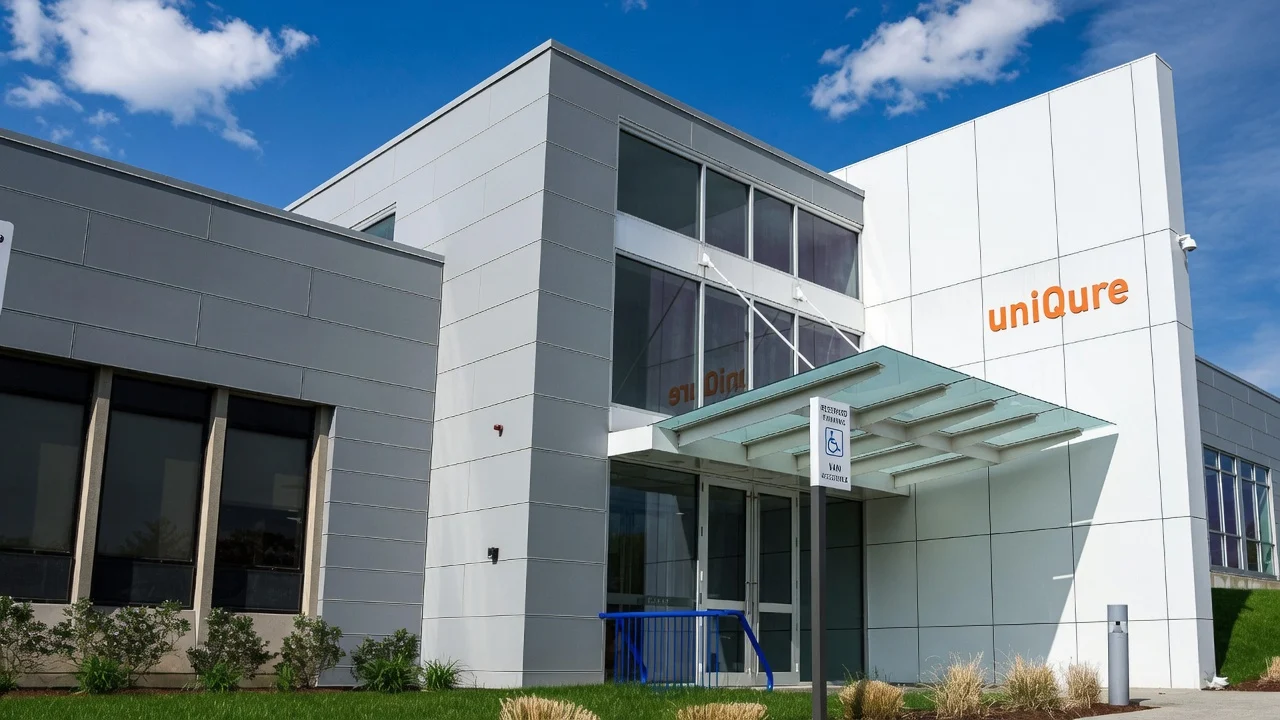
Shares of uniQure (QURE) traded lower by 2.98% to close at $24.09 during Monday’s session, shedding $0.74 in value. The biotechnology company’s stock continued its downward trajectory even as it announced significant progress in the regulatory review of its experimental Huntington’s disease gene therapy. The decline pushed shares beneath the $24.83 technical support threshold as investors processed the company’s advancement through FDA regulatory channels.
Regulatory Agency Accepts uniQure Biologics Application for Huntington’s Therapy
The Food and Drug Administration has accepted uniQure’s Biologics License Application for ifezuntirgene inilparvovec under Priority Review designation. The experimental gene therapy aims to treat Huntington’s disease, a devastating hereditary neurological condition that causes progressive deterioration in movement control, cognitive function, and psychiatric health. The agency designated the submission for Priority Review through its Accelerated Approval mechanism, establishing a target action date of May 1, 2027.
Regulators have signaled their tentative intention to convene an Advisory Committee meeting to evaluate the application, though no firm date has been established for this proceeding. The acceptance represents a crucial advancement toward potential market authorization for the investigational therapeutic candidate.

The submission draws upon 36-month data generated from the company’s Phase I/II clinical development program. Researchers utilized a propensity score-matched external control group derived from the Enroll-HD natural history registry to compare outcomes in treated patients. The company disclosed this comparative analysis publicly on September 24, 2025, establishing the clinical foundation for its regulatory filing.
Confirmatory Study Design Established for 200-Patient Huntington’s Trial
uniQure has reached alignment with FDA regulators on critical design elements for its confirmatory clinical investigation. The planned research will recruit approximately 200 individuals with Huntington’s disease, comparing the gene therapy intervention against standard medical care. Study participants will be randomized at a 2:1 ratio, with twice as many patients receiving active treatment as those continuing standard care alone.
Investigators have selected Total Functional Capacity as the study’s primary efficacy measure at the 36-month timepoint. This assessment tool evaluates patients’ capacity to execute daily living activities and preserve functional autonomy. The company anticipates commencing patient screening activities across U.S. clinical sites during the final months of 2026.
The confirmatory investigation constitutes a critical component of the regulatory framework. Management expects substantial trial progress to occur concurrent with the review period leading to potential product approval. This clinical strategy aims to generate supplementary evidence documenting the therapy’s therapeutic benefit and safety characteristics.
Proprietary Gene-Silencing Platform Earns Multiple FDA Recognitions
Ifezuntirgene inilparvovec—designated internally as AMT-130—employs uniQure’s proprietary miQURE gene-silencing technology platform. The therapeutic approach utilizes an engineered microRNA construct designed to suppress huntingtin gene expression and reduce levels of the potentially neurotoxic exon 1 protein fragment. Administration occurs through a single neurosurgical procedure utilizing MRI guidance to deliver the therapy directly into the striatal region of the brain.
Additionally, AMT-130 has received Breakthrough Therapy designation and Regenerative Medicine Advanced Therapy classification from the FDA. The agency has further assigned Fast Track development status to the program. These multiple regulatory designations underscore the significant agency interest in facilitating development for this rare disease indication.
The company has dosed 51 individuals with AMT-130 throughout its Phase I/II clinical development program. The randomized U.S. investigation enrolled 26 participants, comprising 16 patients receiving active therapy and 10 control subjects undergoing sham surgical procedures. Four individuals initially assigned to the control arm subsequently crossed over to receive treatment following approximately 12 months of observation.
The European open-label investigation included an additional 13 patients distributed across low-dose and high-dose treatment cohorts. A separate group of 12 patients received both dose levels while maintained on immunosuppressive therapy. An additional six U.S. participants are currently evaluating high-dose administration in patients with reduced striatal volumes.
Despite Monday’s Priority Review announcement from the FDA, QURE stock continued experiencing downward pressure throughout the trading session. The breach of the $24.83 support threshold now establishes that level as a significant technical reference point. The May 1, 2027, regulatory target date now represents the next critical catalyst on the company’s developmental timeline.
Get 3 Free Stock Ebooks
Discover top-performing stocks in AI, Crypto, and Technology with expert analysis.
- Top 10 AI Stocks - Leading AI companies
- Top 10 Crypto Stocks - Blockchain leaders
- Top 10 Tech Stocks - Tech giants